+91-8527691232 Paonta Sahib, HP & Noida, India Exporting to 60+ Countries
0%

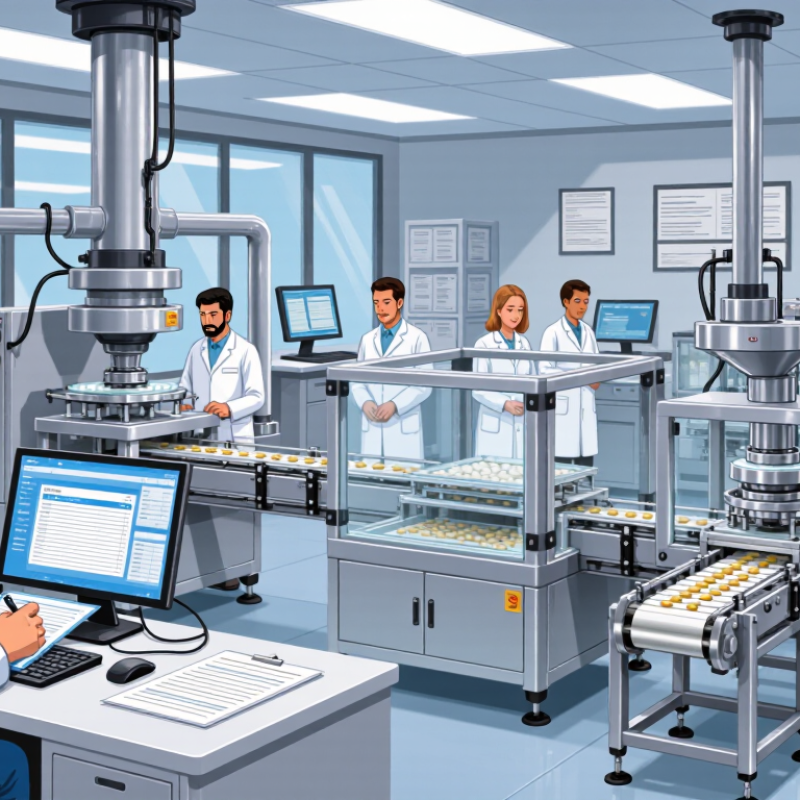
Choosing a tablet or caplet manufacturer is a quality decision, not merely a price comparison. The right partner must protect product performance from formulation through final packaging. In Tablet and Caplet Manufacturing, small process differences can create major commercial problems. A slight compression change may affect hardness, dissolution, or friability. Poor tooling alignment can leave visible defects on the tablet surface.

Dr. Ajaz S. Hussain, a recognized pharmaceutical quality expert, often expresses a central principle this way: “Quality must be built into the product, not tested into it.” This idea should guide every supplier evaluation. Review the manufacturer’s development experience, quality systems, equipment, and technical documentation. Ask how operators monitor blend uniformity, compression force, coating weight, and environmental conditions. Request practical evidence, not attractive claims. Batch records, validation summaries, stability data, deviation reports, and audit responses can reveal far more than a polished website.

Look closely at manufacturing capacity, too. A supplier may produce excellent pilot batches but struggle with consistent commercial volumes. Confirm whether the facility can manage your target batch size, packaging format, and delivery schedule. Check its experience with your active ingredient, excipient profile, tablet shape, and coating requirements. Communication matters during scale-up. Delayed answers often become delayed decisions.

No manufacturer is perfect. I may also overvalue impressive equipment during an initial review. That is a mistake. Skilled people, disciplined procedures, and transparent reporting usually matter more. The best choice combines technical capability, regulatory reliability, realistic capacity, and a willingness to discuss problems openly. This article outlines the questions that separate a dependable partner from a risky supplier.
How to Choose a Tablet and Caplet Manufacturer?

Define the Dosage Form: Tablet or Caplet, Target Dose, and USP <905>

How to Choose a Tablet and Caplet Manufacturer?

Start by defining the dosage form before requesting quotations. A tablet may suit compact, economical dosing, while a caplet can offer a smoother, capsule-like shape for swallowing. The target dose affects tooling, tablet weight, excipient selection, and blend uniformity. Low-dose products need tighter control because small mixing errors can change potency significantly. FDA’s 2023 CDER report recorded 55 novel drug approvals, reflecting continued product diversity and the need for precise dosage planning.

Ask each manufacturer how the proposed formula will meet USP <905>, Uniformity of Dosage Units. The chapter generally uses an acceptance value of no more than 15.0, unless a monograph states otherwise. Request validation data for blend uniformity, tablet weight, hardness, friability, disintegration, and dissolution. FDA’s Process Validation guidance recommends demonstrating consistent commercial-scale performance, not relying only on laboratory results. Examine actual batch records, sampling plans, and out-of-specification investigations. A polished certificate is not enough. I would not treat one passing batch as proof of a capable process. Ask how the manufacturer controls compression force, punch alignment, humidity, and line clearance. These details become visible on the production floor. The right supplier should challenge an unsuitable dose or shape, even when the specification seems acceptable. No simple rule exists.

How to Choose a Tablet and Caplet Manufacturer?

Define the dosage form, target dose, and applicable USP <905> expectations before comparing manufacturing capabilities.

The chart shows key USP <905> reference points for content uniformity: individual dosage units are generally evaluated against 85%–115% of the label claim, while the acceptance value (AV) criterion is commonly set at no more than 15.0. The final specification depends on the product, monograph, and regulatory requirements. A caplet is a capsule-shaped tablet, so the manufacturer should demonstrate suitable tooling, dose accuracy, blend uniformity, and process control for the selected format.

Screen GMP Credentials: FDA 21 CFR Parts 210–211 and WHO GMP Records

How to Choose a Tablet and Caplet Manufacturer?

A manufacturer’s GMP file should be your first practical checkpoint. Ask for evidence, not polished promises. FDA 21 CFR Parts 210–211 address quality systems, production controls, laboratory testing, and recordkeeping. Review whether the site’s procedures match your tablet or caplet process. A certificate alone is insufficient. Check its issuing authority, scope, expiration date, and covered manufacturing activities. Then compare those details with your product requirements.

WHO GMP records can reveal how consistently a facility controls daily operations. Request completed batch records, cleaning logs, equipment calibration reports, and deviation investigations. Look for clear signatures, timestamps, review steps, and traceable changes. Strong records should connect raw-material release with final-product approval. They should also show how rejected materials are segregated and investigated. Do not accept unexplained gaps. A missing page may indicate weak discipline, or simply poor document control. Either possibility deserves a direct question.

During an audit, observe details people often overlook. Are temperature readings recorded near the actual storage area? Do operators follow written instructions without improvising? Ask how recurring deviations become corrective and preventive actions. The answer should include owners, deadlines, effectiveness checks, and supporting evidence. I would also examine recent inspection findings, not only favorable records. That can feel uncomfortable. Yet honest review builds a more reliable supplier decision than attractive paperwork. Consider asking an independent quality professional to challenge your assumptions.

How to Choose a Tablet and Caplet Manufacturer? - Screen GMP Credentials: FDA 21 CFR Parts 210–211 and WHO GMP Records

Evaluation Dimension Evidence to Request Relevant Requirement What to Verify Recommended Screening Outcome
GMP Quality System Current quality manual, organization chart, quality policy, and documented responsibilities FDA 21 CFR 210.10 and 211.22; WHO GMP quality management principles An independent quality unit has authority over approval, rejection, investigation, and release decisions Pass: Responsibilities and release authority are formally documented
FDA Regulatory Status Facility registration details, applicable FDA inspection history, and responses to inspection observations FDA 21 CFR Parts 210–211 Registration or inspection information is current and supported by controlled corrective and preventive actions Note: FDA registration is not the same as FDA approval or GMP certification
WHO GMP Records Recent GMP inspection report, valid certificate where applicable, and corrective action records WHO Technical Report Series No. 986, Annex 2, “Good Manufacturing Practices for Pharmaceutical Products: Main Principles” The scope covers solid oral dosage forms, relevant manufacturing areas, and the intended market Pass: Records are authentic, current, in scope, and consistent with the proposed product
Facility and Equipment Qualification Installation, operational, and performance qualification summaries for critical equipment FDA 21 CFR 211.63, 211.67, and 211.68; WHO GMP premises and equipment requirements Equipment is suitable for tablet or caplet production, cleaned appropriately, maintained, and protected from contamination Pass: Qualification, calibration, cleaning, and maintenance records are complete
Master Manufacturing Documents Master formula, manufacturing instructions, theoretical yield, process parameters, and packaging instructions FDA 21 CFR 211.186 Instructions identify materials, quantities, equipment, processing steps, sampling points, and acceptance criteria Pass: Documents are approved, version-controlled, and product-specific
Batch Production and Control Records Anonymized completed batch record showing weighing, blending, granulation, compression, coating, and packaging entries FDA 21 CFR 211.188 Entries are contemporaneous, attributable, legible, complete, reviewed, and include deviations or discrepancies Pass: No unexplained gaps, overwritten data, or undocumented process changes
In-Process Controls Specifications and results for blend uniformity, tablet weight, hardness, thickness, friability, disintegration, and appearance FDA 21 CFR 211.110 Controls are scientifically justified and performed at defined stages to monitor process consistency Pass: Results meet approved limits and are linked to the batch record
Finished Product Testing Product specification, analytical methods, certificates of analysis, and laboratory review records FDA 21 CFR 211.160, 211.165, and 211.194 Testing covers identity, strength, quality, purity, and relevant performance attributes such as dissolution Pass: Methods are suitable, results are traceable, and an authorized quality unit approves release
Process Validation Validation master plan, process validation protocol, reports, and continued process verification data FDA 21 CFR 211.100(a); FDA process validation guidance; WHO GMP validation principles Critical process parameters and quality attributes are identified, controlled, and supported by documented evidence Pass: Validation reflects the actual product, equipment, batch size, and manufacturing process
Cleaning and Cross-Contamination Control Cleaning procedures, residue limits, cleaning validation reports, and line-clearance checklists FDA 21 CFR 211.42, 211.67, and 211.182; WHO GMP contamination-control principles Controls address active ingredients, excipients, dust, residues, mix-ups, and labeling or packaging clearance Pass: Acceptance limits are justified and cleaning effectiveness is demonstrated
Data Integrity and Electronic Records Data-integrity policy, audit-trail review procedure, access-control matrix, and backup records FDA 21 CFR 211.68(b), 211.180, and 211.194; WHO GMP data-integrity guidance Records are attributable, original or true copies, contemporaneous, accurate, complete, consistent, enduring, and available Pass: User access, audit trails, backups, and record reviews are controlled
Deviation, CAPA, and Change Control Sample deviation investigations, root-cause analyses, CAPA effectiveness checks, and change-control logs FDA 21 CFR 211.100, 211.192, and 211.194; WHO GMP quality system principles Unexpected events are investigated before batch disposition and changes are assessed for product and process impact Pass: Investigations are timely, scientifically justified, and documented to closure
Stability Program Stability protocols, chamber qualification, ongoing stability results, and proposed shelf-life support FDA 21 CFR 211.166 Studies use approved containers and storage conditions and evaluate relevant chemical, physical, and microbiological attributes Pass: Shelf life and storage conditions are supported by documented stability data
Complaints, Recalls, and Traceability Complaint SOP, recall procedure, mock-recall results, distribution records, and product traceability map FDA 21 CFR 211.198 and 211.150; WHO GMP complaints and product-recall principles Each batch can be traced from raw materials through distribution, with procedures for rapid investigation and recall Pass: A documented mock recall demonstrates effective traceability
Record Retention and Availability Record-retention schedule, archival procedure, retrieval test, and controlled document index FDA 21 CFR 211.180 Records are retained for at least one year after the expiration date of the applicable batch, or one year after distribution of the last batch, whichever is longer Pass: Records are secure, protected from alteration, and promptly retrievable
Screening principle: Review original or appropriately redacted records, confirm that the document scope matches the intended tablet or caplet product, and verify that certificates or inspection records are current rather than relying on logos, marketing claims, or registration status alone.

Compare Quality Data: USP <905> Acceptance Value NMT 15.0 and Dissolution

How to Choose a Tablet and Caplet Manufacturer?

When choosing a tablet or caplet manufacturer, ask for measurable quality data. USP <905> evaluates uniformity of dosage units through an Acceptance Value. For many products, an AV of NMT 15.0 is an important batch release criterion. However, this number needs context. Request the individual unit results, calculated mean, variability, and test stage. A passing value alone cannot explain every manufacturing risk.

Dissolution data deserves the same attention. Review the apparatus, medium, volume, agitation speed, sampling times, and acceptance limits. A reliable manufacturer should provide validated methods and clear dissolution profiles. Check whether the method can detect meaningful formulation changes. One fast result may look impressive. It may still hide incomplete release at later time points. Compare several commercial-scale batches, not one carefully selected sample.

I would also examine deviation records, out-of-specification investigations, stability results, and method-transfer evidence. These documents show how the site responds when production is imperfect. That matters. Ask whether operators control compression force, tablet weight, hardness, and coating conditions. Small variations can affect both AV and dissolution. My own concern would be a polished summary without raw data. Strong suppliers explain limitations openly, including borderline results and repeated testing. Good decisions require evidence, not confident language.

Audit Capacity: Batch Size, OOS Trends, Lead Time, and Scale-Up Data

Choosing a tablet and caplet manufacturer requires more than reviewing equipment lists. Audit capacity shows whether the site can manage your product reliably. Request batch records for development, pilot, and commercial lots. Compare planned quantities with actual output, yield, compression speed, and reject rates. Ask whether the proposed batch size matches available blending and compression capacity.

Out-of-specification trends deserve close attention. Review at least 24 months of OOS investigations, including assay, dissolution, hardness, weight variation, and microbiological results. Look for repeated causes, weak root-cause analysis, and overdue corrective actions. A clean dashboard is not evidence. No facility has zero deviations; that may signal poor reporting rather than excellent control. Ask to see investigation timelines and effectiveness checks.

Lead time should include material purchasing, production, laboratory testing, packaging, and batch release. Request the average and longest recent timelines, not only the promised schedule. Scale-up data should connect pilot results with commercial performance. Check process parameters, blend uniformity, compression behavior, and stability observations. During review, I would challenge unexplained gaps between batches. Some explanations may sound reasonable, but missing data still creates risk. A written contingency plan for equipment failure, testing delays, or raw-material shortages can reveal how prepared the manufacturer really is.

Assess Stability and Cost: ICH Q1A 25°C/60% RH and Total Supply Risk

Choosing a tablet or caplet manufacturer requires more than comparing unit prices. Stability evidence should guide the decision. Ask for data generated under ICH Q1A conditions, including 25°C and 60% relative humidity. Review assay, dissolution, hardness, friability, impurities, and appearance at each testing interval.

Look closely at the packaging system. A well-sealed blister may protect moisture-sensitive tablets better than a low-cost bottle. Request the protocol, sampling plan, analytical methods, and trend reports. Do not accept a clean-looking table without understanding its assumptions. Real manufacturing experience matters here. Small changes in compression force or coating thickness can affect dissolution during storage.

Cost also includes total supply risk. Examine active ingredient sources, excipient availability, tooling capacity, utilities, release testing, and transport lanes. Ask whether qualified backup suppliers and alternative production equipment exist. One site may appear efficient, but a shutdown can stop every shipment. That risk is expensive.

Be realistic. No forecast is perfect. Review lead times, minimum order quantities, inventory policies, deviation history, and change-control practices. A manufacturer that explains weaknesses clearly may be more reliable than one presenting flawless figures. Stability data should support confidence, not replace careful questions.

FAQS

How should the dosage form be chosen?

A tablet supports compact, economical dosing. A caplet has a smoother, capsule-like shape. Swallowing comfort matters.

Why is the target dose important?

The dose affects tooling, tablet weight, excipients, and blend uniformity. Low-dose products require tighter mixing control. Small errors matter.

What quality standard should the formula meet?

Ask whether the formula meets USP <905>, Uniformity of Dosage Units. The acceptance value is generally no higher than 15.0.

Which validation records should be reviewed?

Request data for blend uniformity, weight, hardness, friability, disintegration, and dissolution. Examine actual batch records, not polished certificates alone.

What manufacturing controls deserve attention?

Review compression force, punch alignment, humidity, and line clearance. These controls become visible on the production floor. Details matter.

How can manufacturing capacity be audited?

Compare planned quantities with actual output, yield, compression speed, and reject rates. Review development, pilot, and commercial batch records.

Why should out-of-specification investigations be examined?

Review at least 24 months of investigations. Check assay, dissolution, hardness, weight variation, and microbiological results. Repeated causes deserve concern.

What should a realistic lead-time review include?

Include purchasing, production, laboratory testing, packaging, and batch release. Request average and longest recent timelines. Promised schedules may be too optimistic.

How can scale-up readiness be assessed?

Compare pilot results with commercial performance. Check process parameters, blend uniformity, compression behavior, and stability observations. Missing data remains a risk.

What should happen when the proposed dose or shape seems unsuitable?

The manufacturer should challenge the proposal. No simple rule exists. I would question any supplier that agrees too easily.

Conclusion

Choosing a reliable partner for Tablet and Caplet Manufacturing requires more than comparing prices. Start by defining the dosage form, target dose, and applicable USP <905> requirements so the manufacturer can demonstrate consistent dose uniformity. Review its GMP credentials, including compliance with FDA 21 CFR Parts 210–211 and documented WHO GMP practices. Quality data should cover USP <905> acceptance values, with an Acceptance Value of NMT 15.0 where applicable, as well as dissolution performance, analytical methods, and investigation procedures.

A thorough evaluation should also examine production capacity, minimum and maximum batch sizes, out-of-specification trends, lead times, and evidence of successful scale-up. Stability programs should follow ICH Q1A conditions, including 25°C/60% RH when appropriate, to confirm product quality throughout its intended shelf life. Finally, assess total supply risk by considering material sourcing, manufacturing continuity, quality oversight, communication, and long-term cost—not only the initial quotation.
Maris

Maris

Maris is a dedicated marketing professional with a strong understanding of the company’s products, customers, and industry landscape. Through a combination of market research, strategic planning, and clear communication, Maris helps transform complex product information into practical, valuable......